BIA-SOLVE · ENTERPRISE MOLECULAR SIMULATION & AI

Accelerate Molecular Discovery.
Eliminate Costly Wet-Lab Trial & Error.

Specialized atomistic simulations, AI-driven drug design, and QM/MM enzyme engineering for Biotech, Pharma, and Chemical innovators. We turn months of wet-lab screening into targeted in silico decision gates.
University of Coimbra · DEQ / FCTUC / CERES
100% Client IP Ownership & Bilateral NDA
EuroHPC & High-Performance Compute
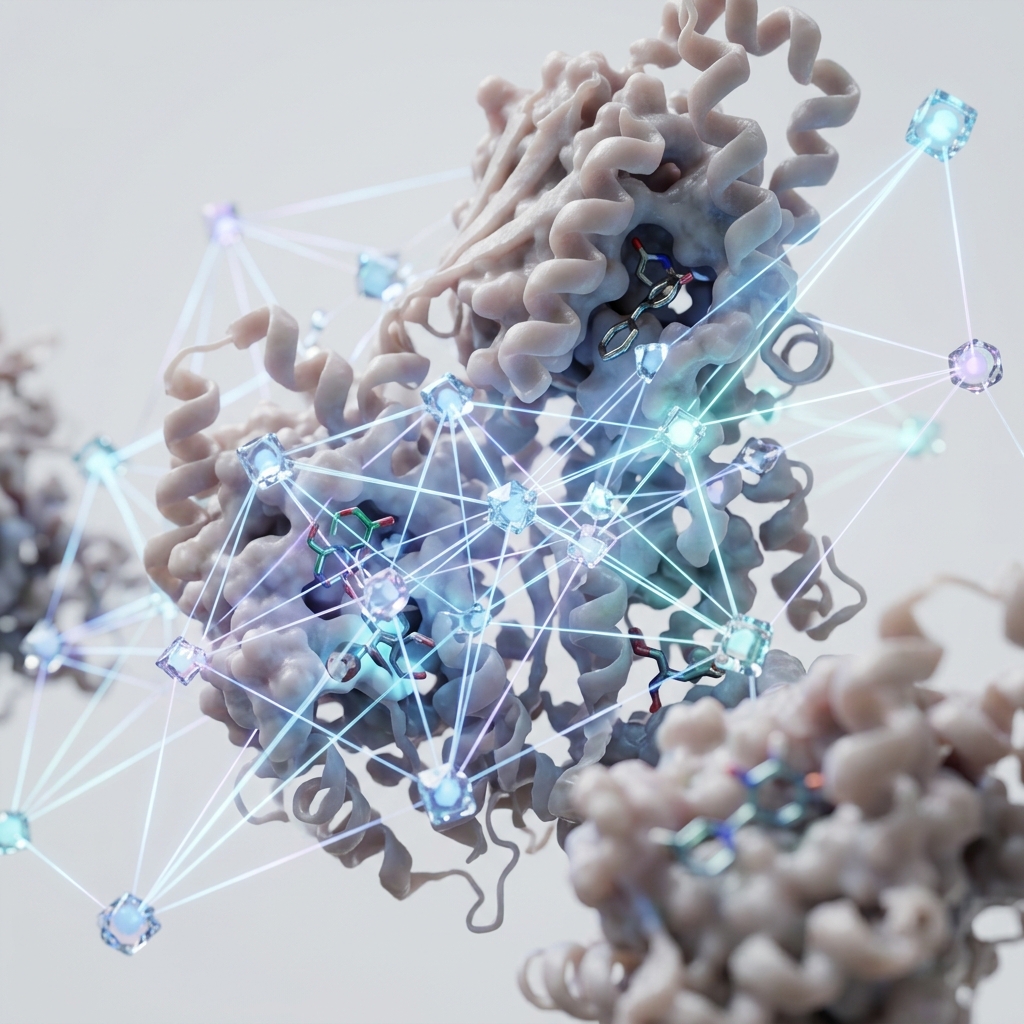
Official BIA Lab 3D Blender Macromolecular Simulation and AI Graph Nodes

100% Client IP Ownership

You own all generated hits, scaffolds, data, and patentable matter.

Strict Bilateral NDA

Executed before any target structures or proprietary data are shared.

Stanford Top 2% Leadership

Led by Dr. Sérgio F. Sousa with 172+ peer-reviewed papers.

Rapid 2–4 Wk Turnaround

Milestone-driven project sprints aligned with your discovery deadlines.
172+
Peer-Reviewed Papers
Published in leading ISI/Scopus scientific journals
5,800+
Citations · h-index 36
World's Top 2% Most-Cited Researchers (Stanford 2022–2024)
€1.0M+
Competitive R&D Grants
CaixaResearch Health & Gilead programs
2–4 Wks
Sprint Turnaround
From computational initiation to actionable wet-lab hit list

Bridging Quantum Physics and Macromolecular Engineering

We leverage advanced 3D visual computing, quantum-mechanical transition state modeling, and atomistic docking to deconstruct complex biological phenomena at single-Angstrom resolution.

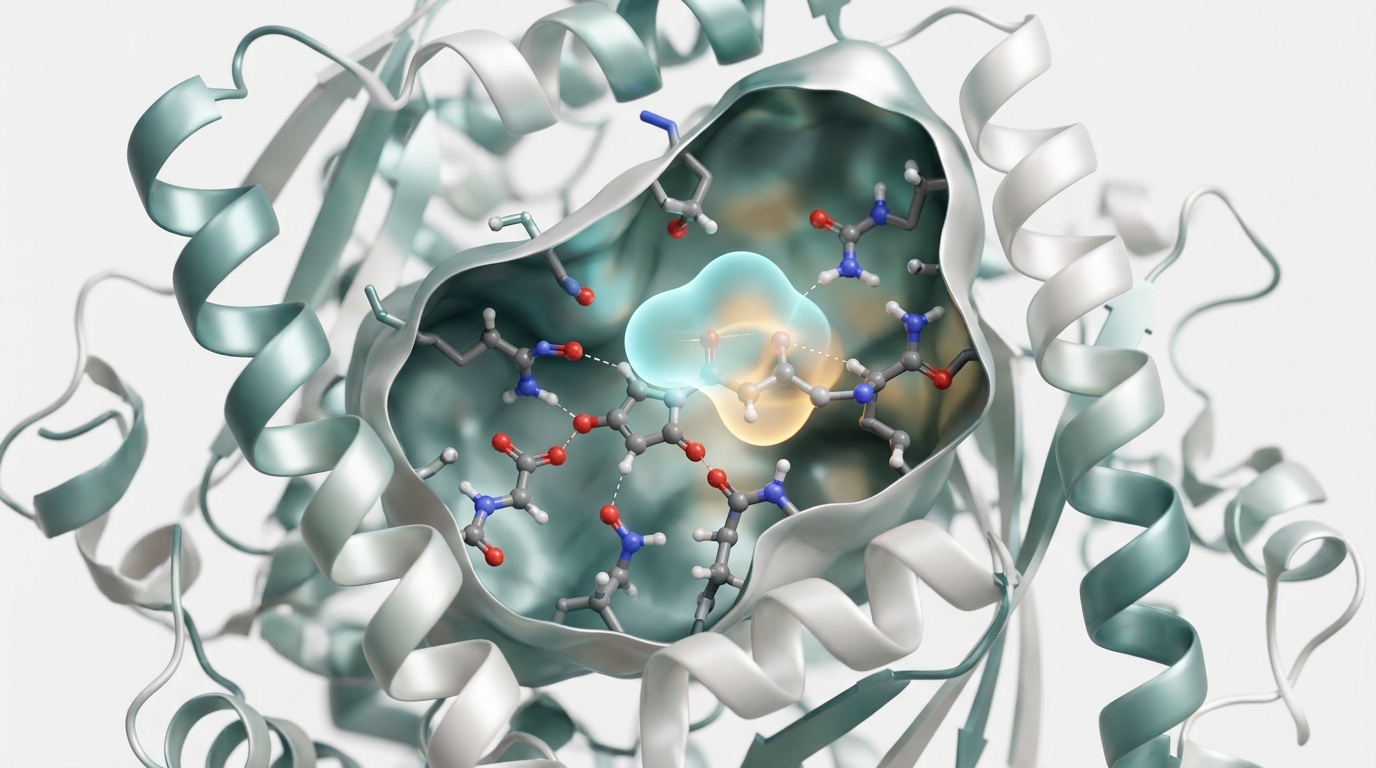
3D Blender Molecular Nodes Render of Enzyme Catalytic Pocket and Substrate Transition State

Rational Enzyme Engineering & Biocatalysis

Unlock sustainable chemical transformations. We employ hybrid Quantum Mechanics / Molecular Mechanics (QM/MM) to characterize catalytic mechanisms and redesign enzyme active sites for novel industrial substrates.

  • Electronic-Level Mechanism: Unambiguous transition state identification and reaction free-energy barriers.
  • Substrate Repurposing: Computational mutagenesis to accommodate non-natural precursors with high turnover.
  • Thermostability Optimization: Engineering robust biocatalysts that endure industrial pH and organic solvents.
Inquire About Biocatalysis →
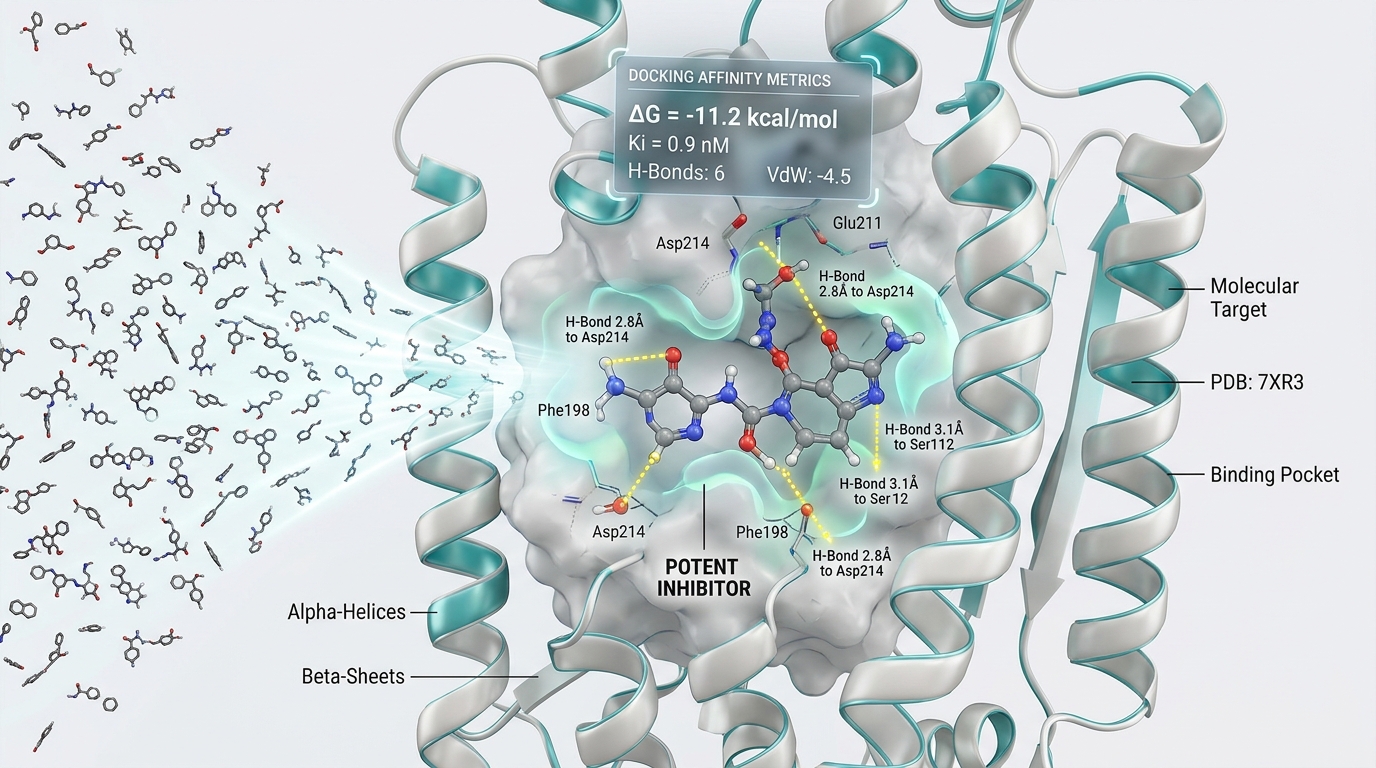
3D Macromolecular Receptor Docking and High-Throughput Virtual Screening

Computer-Aided Drug Discovery (CADD) & Lead Optimization

Eliminate non-viable chemical matter early. Our multi-tiered virtual screening pipelines integrate all-atom molecular docking, consensus scoring, and Free Energy Perturbation (FEP) to prioritize leads with nanomolar potency.

  • High-Throughput Virtual Screening: Mining across 10M+ compound libraries with multi-conformation scoring.
  • Thermodynamic Binding Kinetics: Free energy calculations and allosteric residence time predictions.
  • Predictive ADMET Profiling: Early identification of metabolic stability, solubility, and safety profiles.
Inquire About Drug Discovery →

Core Computational Services for Industry

Rooted in over two decades of peer-reviewed computational excellence at the University of Coimbra, our services provide actionable, investor-grade scientific data.

01 // PHARMA

High-Throughput Virtual Screening

Screen millions of compounds against your validated protein targets using consensus docking algorithms and strict pharmacokinetic filters.

  • Structure-based & ligand-based virtual screening
  • Consensus scoring & receptor flexibility docking
  • Lead optimization & scaffold hopping analysis
BUSINESS IMPACT Cut discovery turnaround from months to weeks; eliminate non-viable synthesis costs.
02 // BIOPHYSICS

Molecular Dynamics & Kinetics

Quantify conformational stability, allosteric pathways, and ligand residence times across microsecond all-atom physiological simulations.

  • Microsecond all-atom & coarse-grained simulations
  • Binding kinetics and residence time estimations
  • Membrane permeability & protein-lipid interactions
BUSINESS IMPACT Ensure biological efficacy and target engagement prior to in vitro validation.
03 // BIOCATALYSIS

Enzyme Engineering & QM/MM

Direct transition-state modeling to guide rational enzyme mutations, boost turnover numbers (kcat), and enable green synthetic routes.

  • Full QM/MM electronic reaction barrier calculations
  • Active site redesign for non-native substrates
  • Thermal stability and solvent tolerance engineering
BUSINESS IMPACT Replace hazardous chemical catalysts with clean, proprietary biocatalysts.
04 // AI / ML

AI & Generative Molecular Design

Harness deep generative architectures and predictive machine learning models to invent novel, synthetically tractable chemical entities.

  • AI-driven de novo molecular generation & optimization
  • Predictive QSAR & physicochemical property algorithms
  • AI protein structure prediction (AlphaFold/esmFold)
BUSINESS IMPACT Discover patentable chemical scaffolds far outside competitor IP territory.
05 // DEGRADERS

Targeted Protein Degradation & PROTACs

Rational in silico design of bifunctional degraders (PROTACs) and molecular glues to selectively eliminate disease-causing targets via ubiquitin-proteasome recruitment.

  • Ternary complex modeling (Target · PROTAC · E3 Ligase)
  • Linker flexibility, length & conformational sampling
  • Cooperativity analysis & protein-protein interface scoring
  • Anti-target docking & selectivity screening against off-target risks
BUSINESS IMPACT Tackle historically "undruggable" proteome targets, de-risking bifunctional synthesis before wet-lab investment.
06 // R&D GRANTS

Consortia & Scientific Advisory

Flexible collaboration formats: tactical discovery sprints, dedicated corporate advisory, or joint grant co-applicants (Horizon Europe).

  • Rapid feasibility audits and target de-risking reports
  • Horizon Europe & national R&D grant co-application
  • Executive scientific dossiers for VC and patent filings
BUSINESS IMPACT Access tier-1 university scientific infrastructure with zero internal overhead.

Solving What Stalls Early-Stage Pipelines

Biotech and pharma programs rarely fail from lack of ambition—they stall on molecular bottlenecks. Here is how our in silico strategies eliminate high-risk wet-lab roadblocks.

R&D Roadblock SBDD & CADD

Challenging & "Undruggable" Targets

Your target lacks rigid, well-defined binding pockets, features high conformational flexibility, or relies on shallow protein-protein interaction (PPI) interfaces that defeat classical docking.

Our In Silico Strategy

All-atom conformational ensemble sampling, cosolvent molecular dynamics for cryptic pocket detection, and multi-conformation consensus screening to locate druggable transient states.

Outcome: Unlocks validated hit-finding starting points for previously refractory targets.
Safety Liability Lead Optimization

Off-Target Toxicity & Selectivity Traps

Potent hit molecules frequently cross-react with homologous kinase isoforms, GPCRs, or anti-targets (hERG, CYP-450), threatening to derail in vivo efficacy or clinical safety.

Our In Silico Strategy

Counter-screening docking panels and our Gulbenkian Award-winning Anti-Target profiling methodology to map selectivity margins and redesign compounds with micro-substitutions.

Outcome: Rescues promising chemical series by engineering clean isoform selectivity.
Modality Complexity PROTACs & Degraders

PROTAC Ternary Instability & Linker Guesswork

Bifunctional degraders require productive ternary complex formation (Target · PROTAC · E3 Ligase). Synthesizing dozens of arbitrary linker lengths and attachments is prohibitively expensive.

Our In Silico Strategy

Full ternary complex sampling, protein-protein interface cooperativity calculation, and conformational linker optimization grounded in our €1M CaixaResearch Health degrader program.

Outcome: Focuses synthetic chemistry strictly on ternary-productive degrader architectures.
Bioprocess Hurdle Enzyme Engineering

Enzyme Inactivation & Poor Enantioselectivity

Wild-type biocatalysts often lose catalytic activity at industrial temperatures, denature in organic solvents, or fail to yield enantiomerically pure pharmaceutical intermediates.

Our In Silico Strategy

Hybrid QM/MM transition state mapping to compute activation barriers, combined with rational active site mutagenesis and surface engineering for thermal and solvent durability.

Outcome: Replaces toxic transition-metal chemistry with highly selective, robust biocatalysts.

What You Receive in Hand

We deliver actionable, experiment-ready chemical intelligence—not raw data dumps. Every project terminates in a turnkey package designed for immediate laboratory ordering and wet-lab testing.

Ranked Candidate Hit Matrix

  • Multi-parametric scored and filtered hit list.
  • Commercial vendor catalog IDs (Enamine, Mcule, Chemspace) ready for one-click purchasing.
  • Predicted physicochemical, ADMET, and PAINS clearance.

Clean Atomic Coordinates

  • Complete 3D coordinates in standardized PDB, SDF, and Mol2 formats.
  • All protonation states, tautomers, and residue partial charges parameterized.
  • Equilibrated receptor-ligand and ternary complex models.

Thermodynamic & Free Energy Dossier

  • Computed binding free energies (ΔG) and affinity correlations.
  • Catalytic activation barrier profiles (ΔG‡) for enzyme reactions.
  • Molecular dynamics RMSD/RMSF and residence time estimations.

3D Molecular Trajectories & Visuals

  • Publication-ready 3D structural figures and pocket binding poses.
  • Atomistic simulation video trajectories capturing allosteric events.
  • Interactive PyMOL/ChimeraX sessions for internal team review.

Executive Decision Deck

  • Comprehensive PowerPoint / PDF summary built for R&D committees.
  • Clear Go / No-Go experimental recommendations and assay selection.
  • Intellectual property defense dossiers for patent applications.

Direct Scientific Debriefing

  • 1-on-1 virtual consultation with Group Leader Dr. Sérgio F. Sousa.
  • Live interactive walkthrough of 3D binding poses and mechanisms.
  • Collaborative Q&A to align in silico predictions with your wet-lab biology.

Built for Your Organization's Milestones

Whether you are an emerging venture pitching to venture capital or an established multinational optimizing API yield, our workflows fit seamlessly.

BIOTECH & VENTURES

Biotech Startups & Spin-offs

Secure investor validation with robust in silico target engagement and hit identification without the multi-million capital expense of an in-house HPC lab.

Seed / Series A Rapid Milestones IP Protection
PHARMACEUTICALS

Pharma & Drug Developers

Augment your medicinal chemistry departments with high-throughput screening, free energy perturbation (FEP) pipelines, and ADMET liability mitigation.

Lead Optimization Binding Kinetics De-risking
GREEN INDUSTRY

Chemical & Biocatalysis

Engineer proprietary enzymes for bio-based materials, sustainable specialty chemicals, flavorings, and active cosmetic ingredient synthesis.

Green Chemistry Enzyme Design Yield Boost
ACADEMIA & HUBS

Research Centers & CoLabs

Fortify international consortia, co-patents, and competitive grant proposals (Horizon Europe) with world-class computational chemistry leadership.

Horizon Europe Joint Patents High Impact

Transparent Engagement Models

We offer clear, contractually flexible collaboration frameworks designed to fit startups, global pharmaceutical enterprises, and academic-industrial consortia.

Dedicated In Silico FTE
Embedded Retainer · Continuous Support

An embedded computational chemistry extension to your internal medicinal chemistry team, supporting recurring iterative Design-Make-Test-Analyze (DMTA) cycles.

  • Dedicated senior computational modeling bandwidth.
  • Continuous analog profiling, docking, and SAR exploration.
  • Weekly technical meetings and real-time project steering.
  • Priority allocation on dedicated HPC clusters.
Consortia & Grant Co-Dev
Co-Funded R&D · Horizon Europe

Joint scientific leadership for collaborative European and international grants, leveraging advanced computational infrastructure and university co-funding mechanisms.

  • Co-applicant for Horizon Europe (Pillar II Health & Bio).
  • Shared risk and co-development intellectual frameworks.
  • Access to high-performance computing (HPC) resources.
  • High-impact joint patents and tier-1 publications.

From First Briefing to Decision-Ready Results

A structured, milestone-driven framework designed to deliver validated computational insights within your R&D deadlines, under strict confidentiality.

01

Scoping & Confidentiality

We establish a bilateral NDA, review your target's structural data and feasibility, and define precise project milestones and deliverables.

02

Computational Execution

We deploy validated molecular simulation, quantum chemistry, and machine learning workflows on dedicated high-performance computing clusters.

03

Decision-Ready Deliverables

You receive prioritized candidate lists, structural rationales, and full coordinate data, followed by a direct scientific debriefing to guide experimental testing.

Discuss Your Molecular Project Directly

Connect with our group leader at the University of Coimbra. We will evaluate your targets under strict confidentiality and propose an optimized computational roadmap.

Dr. Sérgio F. Sousa

Group Leader · BIA Lab
Dept. of Chemical Engineering · University of Coimbra
DEQ · Polo II · University of Coimbra · 3030-790 Coimbra, Portugal

Direct Project Briefing

Specify your target challenge below to generate a pre-structured consultation briefing.